Medical Writing

Regulatory submission scope

We provide medical document writing, structural organisation and consistency review. The client is responsible for signing and submitting documents.

For regulatory submission requests, we confirm engagement boundaries before quoting. The scope below is divided into Can take on, Recommended as a joint engagement and Not taken on.

01Medical Writing

Documents and engagement status

Can take on: 5 items; Recommended as a joint engagement: 5 items; Not taken on: 3 items.

Can take on5
  • Clinical study reports (CSR)

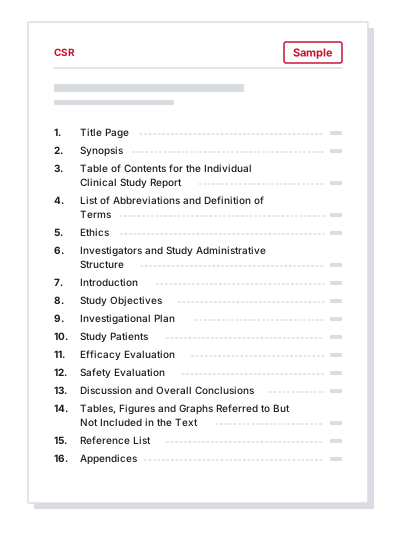
    Written to the structure of chapters 1–16 of the International Council for Harmonisation of Technical Requirements for Pharmaceuticals for Human Use (ICH) E3.

  • Investigator's brochure (IB) updates

    Annual updates and incorporation of safety information based on an existing version.

  • Common technical document (CTD) 2.7 clinical summary

    Clinical summaries organised to the structural requirements of ICH M4E(R2).

  • Regulatory meeting briefing packages

    Writing and compiling meeting application materials; attendance by our staff is not included.

  • CTD 2.6 nonclinical summaries and nonclinical study report writing support

    Writing based on study reports and data supplied by the client, without conducting additional experiments; nonclinical study reports are reviewed and signed by the study director at the research institution.

Recommended as a joint engagement5
  • Lead writing of a clinical trial protocol for a first investigational new drug (IND) application

    Completed jointly with the client's regulatory or medical department, or a partner with a comparable track record.

  • CTD 2.5 clinical overview

    Requires critical analysis of the overall clinical development programme; joint engagement is recommended.

  • Investigator's brochure (IB): first edition

    Requires synthesis of nonclinical and early clinical materials into an overall assessment; joint work with the client's nonclinical and medical departments is recommended.

  • CTD 2.4 nonclinical overview

    Requires an overall critical analysis of nonclinical studies; a joint engagement is recommended.

  • Supporting drafts for responses to regulatory questions

    Responses have tight deadlines and must align with the overall submission strategy; joint work with the client's regulatory department or a partner with relevant experience is recommended.

Not taken on3
  • Electronic Common Technical Document (eCTD) publishing and submission

    Outside the scope of this writing service.

  • Formal communication with regulatory review agencies and in-person defence

    Our professionals all work overseas; we do not undertake these activities.

  • Design and conduct of experimental studies, and GLP compliance responsibility

    We do not conduct experimental (laboratory or animal) studies. GLP compliance responsibility rests with the research institution.

02Medical Writing

Engagement arrangements and responsibilities

For a first engagement, we recommend starting with one document or chapter, reviewed and signed by the client's regulatory or medical department.

We may also undertake the writing component as a consortium member or subcontractor to the lead contract research organisation (CRO). We do not act as the entity responsible for the regulatory application.

Applicant

Article nine of the Measures for the Administration of Drug Registration (药品注册管理办法) (State Administration for Market Regulation Order No. 27, 2020) requires the applicant to be an enterprise or drug development institution able to assume the relevant legal responsibilities. We undertake the agreed document work; the client is responsible for signing and submission.
Sponsor's ultimate responsibility

ICH E6(R3), Annex 1, section 3.6.6 allows sponsors to transfer trial-related activities in accordance with applicable regulatory requirements, while retaining ultimate responsibility for their trial-related activities. Our contracts follow this allocation.

Source verification

Article forty-five of the Measures for the Administration of Drug Registration (药品注册管理办法) provides for extended inspections of other entrusted institutions where necessary. We deliver evidence traceability tables and quality control logs, retaining the source and verification date for each claim.

03Medical Writing

Responsibilities for specialist documents

Drug safety

We write the agreed periodic reports and safety documents. We do not collect, assess or submit individual case safety reports, or act as the entity responsible for pharmacovigilance.

Health economics

We can deliver working health economic model files in Excel or R, as specified by the client, and write the economic sections of a value dossier.

Medical device clinical evaluation

We undertake clinical evaluation reports (CER), post-market clinical follow-up (PMCF) documents and clinical evaluation materials for registration in China. CER evaluators are appointed by the client and sign the evaluation report; we conduct evidence searches, appraisal and writing.

These specialist materials do not change the regulatory submission engagement status listed on this page. The client remains responsible for signing and submitting documents.

04Medical Writing

Regulatory submission track record

Regulatory submissions are a service area we are developing. We do not yet have a prior track record that we can cite publicly for NMPA regulatory submission projects.

If a project requires suppliers to provide a comparable regulatory submission track record, the client should state this when requesting a quotation. Team academic roles, records of other services and public research examples do not substitute for that track record.

A first engagement may be limited to one document or chapter. Further collaboration can be agreed after the first round, once both parties have established how they will work together.

05Medical Writing

Information needed to confirm scope

Please provide document types, the current stage, deadlines and the status of available materials, and state whether joint engagement, supplier onboarding documents or a comparable regulatory submission track record are required. We confirm engagement boundaries before issuing a requirement confirmation form and itemised quotation.

Clinical study report contents sample
Figure 2Clinical study report contents sampleIllustrative sample
Figure caption

Shows the chapter organisation of a CSR for comparison with the document structure.

Get started · Submit a request

Submit a medical writing request

Please provide the document type, quantity, project stage, intended use, available materials and deadline requirements. We first issue a requirement confirmation form, followed by an itemised quote and schedule.

Submit a medical writing requestReply within 24 hours (business days first) · contact@yantohealth.cn